A cleanroom, an ISO 13485 quality system, and a broad list of processes do not, by themselves, demonstrate readiness for Class III medical device contract manufacturing. The relevant question is whether the manufacturer’s controls and records support your device’s specific requirements and can withstand regulatory scrutiny.
A contract manufacturer can contribute substantial quality and process expertise, but device classification is not a manufacturing certification. Engaging a CDMO does not automatically transfer the device company’s regulatory responsibilities. Before production begins, define who owns each activity, what validation evidence is required, and which records must be maintained and reviewed.
This article explains how classification can influence manufacturing planning and what to assess when selecting a partner. It covers process controls, validation, quality responsibilities, documentation, and a controlled transfer into production. It also distinguishes demonstrated capabilities, such as precision machining, laser processing, assembly, testing, and packaging, from general supplier claims, so you can ask for evidence that matches your program.
Key Takeaways
- Class III classification depends on intended use and device type. Verify the applicable regulatory pathway for the specific device.
- Assess Class III medical device contract manufacturing using documented, device-relevant evidence, not broad claims about experience or equipment.
- Distinguish the FDA’s PMA pathway from 510(k), and check pathway details and exceptions against device-specific FDA guidance.
- Plan production transfer in stages: define requirements, transfer processes, validate controls, and approve production readiness using an approach suited to process risk.
- Compare a potential partner’s relevant capabilities, such as machining, laser processing, assembly, testing, and packaging, with your program’s needs.
What Class III Medical Device Contract Manufacturing Involves
Class III medical device contract manufacturing is a defined set of manufacturing activities performed by a contract manufacturer for a device classified by the FDA as Class III. Classification describes a device’s regulatory risk category. It does not certify a factory or establish that a manufacturer is authorized to produce every Class III device. The manufacturer’s work and supporting evidence are determined by the customer agreement and quality arrangements.
How FDA classifies Class III medical devices
The FDA uses a risk-based system with three classes, reflecting the level of regulatory control needed to provide reasonable assurance of safety and effectiveness. A foundational overview of the three classes of medical devices explains this framework. General controls apply across device classes. Special controls address particular risks for some devices, generally within Class II. Class III devices are subject to general controls and typically require premarket approval (PMA), subject to device-specific exceptions.
Classification depends on intended use and the device’s specific characteristics, not just its broad category. Before planning production, check the FDA Product Classification Database for the relevant product code and regulatory requirements, then confirm the pathway for the device. A label such as “implant” or “diagnostic device” alone does not establish its classification.
What a contract manufacturer contributes
A contract manufacturer’s scope is set by the customer agreement. Depending on the program, it may include engineering transfer, process development, assembly, testing, packaging, or production. The agreement and quality documentation should identify which party performs and approves each activity, controls changes, maintains records, and addresses nonconforming product. Outsourcing manufacturing work does not, by itself, transfer the customer’s regulatory responsibilities.
For example, a partner might receive approved design inputs, develop a controlled assembly process, document testing, and package finished devices to agreed specifications. Each activity needs defined acceptance criteria and records. The useful question is not simply whether the partner has certain equipment, but whether it can connect your requirements to repeatable processes and reviewable evidence. Explore the broader range of medical device contract manufacturing services when defining the activities your program may require.
Fong’s reports capabilities in precision machining, laser processing, cleanroom assembly, testing, and packaging. These may be relevant to a project, but capability alone does not establish device-specific readiness or marketing authorization. Confirm which processes apply, what validation evidence is available, and how responsibilities will be documented before agreeing on the production scope.
How Class III Regulatory Pathways Shape Manufacturing Requirements
A regulatory pathway can shape more than the submission file. It helps define the manufacturing evidence a device owner may need, from approved specifications and process records to traceability and supplier oversight. For Class III medical device contract manufacturing, the customer and CDMO should translate program expectations into a clear, documented scope rather than assume one standard set of controls fits every device.
Confirm the device classification and applicable pathway
Many Class III devices require FDA Premarket Approval (PMA), a pathway that evaluates evidence of safety and effectiveness. FDA 510(k) is distinct: it generally focuses on demonstrating substantial equivalence to a legally marketed predicate device. Do not treat 510(k) as the default for Class III. The FDA explains the Premarket Approval (PMA) process, while device-specific exceptions and requirements must be checked individually.
Verify classification and the submission route for the specific device by confirming its FDA product code, class, intended use, indications for use, device characteristics, and applicable regulations. Use FDA classification resources as a starting point, then involve qualified regulatory professionals for pathway questions. A CDMO can provide manufacturing input, but should not be treated as legal counsel or the owner of the customer’s regulatory strategy.
Translate regulatory expectations into manufacturing controls
Once the pathway and program requirements are understood, document which party owns each manufacturing activity and record. Specify what the CDMO must maintain, how records will be reviewed, and how changes will be assessed and approved. This turns broad expectations into practical controls and makes responsibilities visible across the supplier relationship.
- Specifications: Identify the current, approved drawings, materials, work instructions, and acceptance criteria that govern production.
- Process records: Define the evidence to retain for production, inspection, testing, and any required process validation.
- Traceability: Set requirements for linking materials, components, production steps, and test results to the relevant device or production record.
- Change control: Agree how proposed changes are documented, evaluated for impact, and authorized before implementation.
Validation provides documented evidence that a process can consistently achieve its intended result. The appropriate approach depends on the process and its risks. Controlled change review helps preserve that evidence when equipment, materials, methods, or specifications change. Together, these controls connect regulatory expectations to day-to-day production and form part of broader medical device manufacturing compliance planning.
For a program-specific discussion of manufacturing scope and required evidence, contact Fong’s about your device requirements.
How to Evaluate a Class III Medical Device Contract Manufacturer
When evaluating Class III medical device contract manufacturing, look for evidence behind each capability. Advanced equipment does not establish readiness on its own. The manufacturer’s processes, controls, and records must match your device’s materials, tolerances, assembly sequence, testing requirements, and production needs.
Compare suppliers against the same criteria. Request examples and records relevant to your program, with confidential information appropriately protected.
| Evaluation area | Evidence to review |
|---|---|
| Quality system | Applicable quality-system scope, document controls, supplier oversight, and responsibility interfaces. |
| Process capability | Equipment, operator methods, and inspection plans mapped to specifications, materials, and tolerances. |
| Validation | Relevant protocols, acceptance criteria, results, and records showing how process performance is assessed. |
| Traceability | Examples showing how required materials, production steps, inspections, and test results are recorded and linked. |
| Scale-up | A proposed approach for maintaining process controls and acceptance criteria as production requirements change. |
Which technical capabilities should match the device?
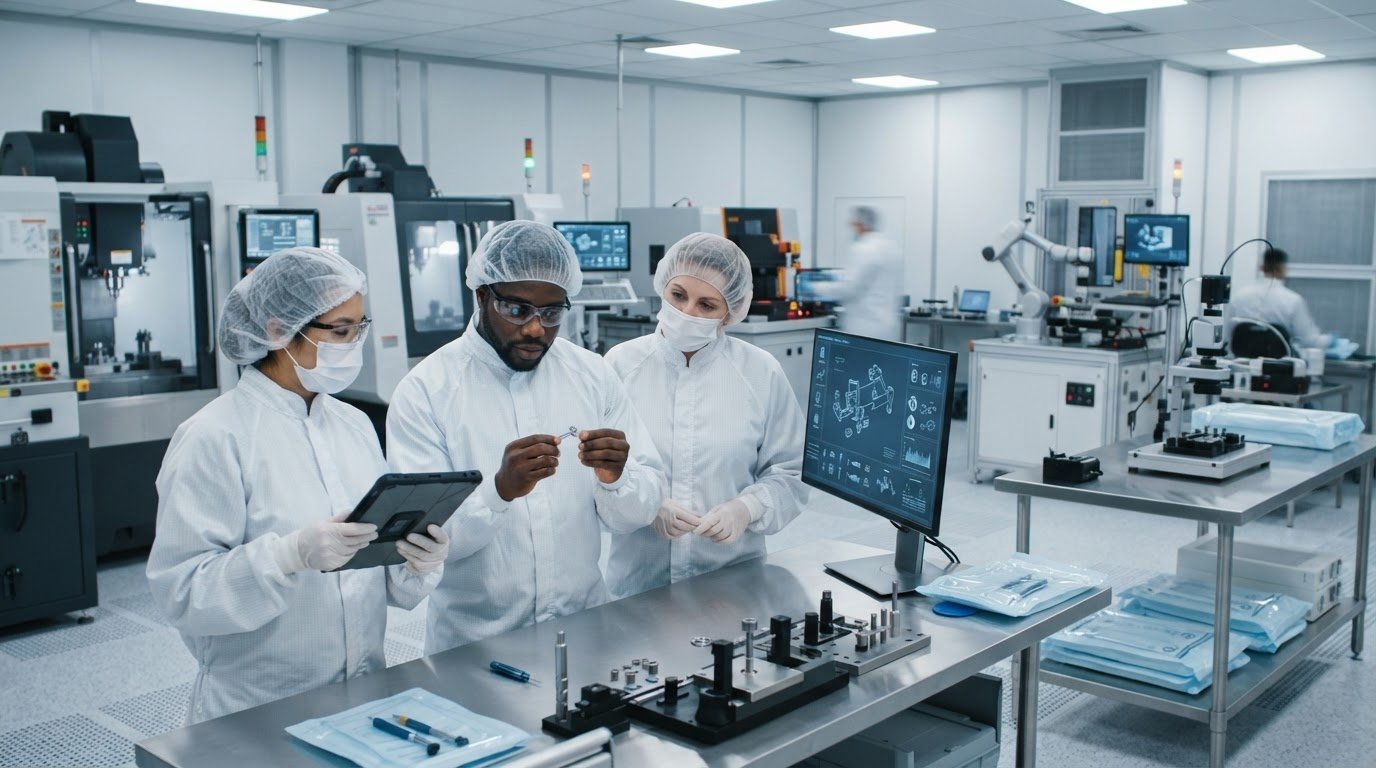
Map each required operation to the design. This could mean precision machining or laser processing for specified features, molding where the design calls for it, and assembly and testing that follow the product’s acceptance criteria. Ask how equipment capability connects to operator instructions and inspection methods. A Class 8 cleanroom is relevant when the product and process require controlled assembly; the classification alone does not establish suitability for a particular device.
Fong’s reports capabilities including precision machining, laser processing, cleanroom assembly, and testing. Treat these as potential areas of fit, then confirm which apply to your device and what supporting evidence is available. Its reported ISO 13485 quality systems should also be assessed for relevant scope, sites, and processes.
What quality and validation evidence should buyers review?
Request program-relevant validation protocols and completed records, including acceptance criteria and documented handling of deviations. Where applicable, review how risk management, CAPA, PFMEA, and change control connect between your organization and the supplier. Design transfer should preserve critical requirements while confirming that selected processes can meet them. Medical device design for manufacturability can help inform process fit, but it does not replace verification against approved specifications.
Supplier controls are part of qualification. The Cornell Legal Information Institute’s Purchasing controls page provides context on supplier evaluation under the former QSR. Verify current obligations against applicable requirements, including FDA’s QMSR.
- Which records will you maintain, and how will we review them?
- How are process changes assessed and approved?
- What evidence demonstrates capability for our specifications and production needs?
How to Plan Validation, Traceability, and Production Transfer
A controlled transfer turns an approved design and agreed requirements into repeatable production. For Class III medical device contract manufacturing, align the customer and CDMO on responsibilities, records, validation evidence, and change handling before routine production begins. Use a staged plan:
- Define requirements. Confirm current specifications, test methods, sampling plans, acceptance criteria, and required production records.
- Transfer processes. Provide controlled drawings, materials information, work instructions, equipment requirements, and relevant process knowledge. Record open risks and assign owners.
- Validate controls. Use documented protocols and results to show that applicable processes can perform as intended under defined conditions.
- Approve production readiness. Review transfer and validation evidence, resolve open issues, and document who authorizes routine production.
Set responsibilities and acceptance criteria before transfer
Document who holds design authority, who owns each production process, and who approves specifications, validation results, and changes. The quality agreement or equivalent controlled documentation should also define supplier controls, access to records, nonconformance escalation, and communication routes between teams. Agree on test methods and acceptance criteria before validation begins, so both parties have a consistent basis for deciding whether the process is ready.
Use validation and production evidence to manage change
IQ, OQ, and PQ are common validation stages. Installation Qualification checks whether equipment is installed as specified. Operational Qualification evaluates whether it operates within defined limits. Performance Qualification assesses whether the process produces acceptable results under intended production conditions. The appropriate approach depends on process risk and should be justified for the specific process, rather than applied as a fixed template.
Maintain traceability appropriate to the device and agreed requirements. Records may link device or lot identifiers to materials, production steps, inspections, and test results. Define how nonconforming product is identified, contained, investigated, and dispositioned. These records connect production outcomes to decisions and follow-up actions.
Document changes to equipment, materials, methods, software, or specifications and assess their impact before implementation. Define who reviews and approves a change, whether customer notification is required, and when additional validation or revalidation is warranted. For complex medical device assembly, clear assembly instructions and in-process testing can be included in the transfer plan where they fit the device and process.
Discuss your manufacturing requirements with Fong’s to review potential process scope and required evidence.
Why Consider Fong’s for Class III Device Manufacturing Support
A manufacturing partner can contribute process expertise and production evidence, but it does not replace the customer’s regulatory strategy or assume regulatory responsibilities by default. For Class III medical device contract manufacturing, assess Fong’s against the specific work your program requires. Confirm responsibilities, evidence, and quality-system scope before defining an engagement.
Where Fong’s capabilities may fit a device program
Fong’s reports medical device CDMO and OEM capabilities that include precision machining, laser processing, cleanroom assembly, product testing, and packaging. These may suit programs requiring controlled component production, laser-processed features, assembly operations, or defined testing and packaging steps. Fit depends on the device’s approved specifications, materials, tolerances, process risks, and acceptance criteria, not simply on the presence of a particular technology.
Fong’s also reports Class 8 cleanroom facilities for disposable and reusable device assembly. A cleanroom classification describes a facility capability, not proof that the environment is suitable for every Class III device or process. Confirm the environmental requirements for your product and the controls relevant to the proposed assembly work.
Smart manufacturing capabilities include real-time monitoring and in-process testing. These can support production visibility and process checks, but they do not replace device-specific validation or demonstrate regulatory approval. Fong’s lists IQ/OQ/PQ validation frameworks and validation support. Discuss which processes apply and request evidence relevant to your program. The company also reports ISO 13485 quality systems. Confirm the applicable scope, sites, and processes directly rather than assuming every activity or program is covered.
What to include in an initial manufacturing discussion
A focused first discussion helps both parties assess fit. Prepare the device specifications, intended manufacturing scope, required processes, and current development stage. Identify the evidence and deliverables your quality and regulatory teams expect, including validation documentation, inspection and test records, and packaging requirements. If sterilization is part of the program, clarify the requirement and responsibility rather than assuming it is included in the manufacturing scope.
- Which proposed processes and facilities are relevant to the device requirements?
- What quality-system scope, validation records, and project-specific evidence can be reviewed?
- How will customer and CDMO responsibilities, approvals, and changes be documented?
Use the answers to establish whether the proposed scope is supported by relevant capability and documentation. Share your device requirements, discuss the manufacturing scope, and contact Fong’s to discuss your project.
Move Your Manufacturing Plan Toward Readiness
Strong Class III medical device contract manufacturing depends on more than access to equipment. Confirm the device-specific regulatory pathway, define customer and CDMO responsibilities, and assess whether process controls, validation records, and traceability requirements match your program.
Fong’s reports ISO 13485 quality systems and documented IQ/OQ/PQ validation frameworks, with applicable scope to verify. Its capabilities include Class 8 cleanroom assembly, advanced laser processing, and smart manufacturing with real-time monitoring and in-process testing. These may support relevant manufacturing needs, but they do not replace your regulatory strategy or establish suitability without a program-specific review.
Bring your device specifications, intended manufacturing scope, process requirements, and expected documentation to an initial discussion. Together, you can identify potential capability fit and the evidence and responsibilities that need confirmation. Discuss your Class III manufacturing requirements with Fong’s.
A disciplined evaluation creates a clearer path from design transfer to controlled production. Aligning requirements and evidence helps your team make informed decisions about moving forward.
Frequently Asked Questions
Does every Class III medical device require FDA premarket approval?
No. Many Class III devices follow the FDA Premarket Approval (PMA) pathway, but requirements depend on the specific device, its classification, and applicable regulations. A broad product description is not enough to determine the route. Confirm the device’s FDA product code and current requirements, and obtain appropriate regulatory advice. A contract manufacturer can support agreed manufacturing controls and records, but cannot guarantee FDA authorization or determine the device company’s full regulatory obligations.
What does a Class III medical device contract manufacturer do?
A contract manufacturer performs the development or production activities defined in its agreement with the device company. The scope may include process engineering, component manufacturing, assembly, testing, packaging, and validation support. The parties should document who owns design decisions, quality responsibilities, records, and regulatory submissions. The device company remains responsible for confirming its regulatory strategy and ensuring the overall program meets its obligations. A clear agreement defines the manufacturer’s role and reduces ambiguity.
Can a CDMO manufacture a Class III medical device?
Yes. A CDMO may support Class III device manufacturing if its relevant processes, quality systems, facilities, and personnel fit the program requirements. Buyers should request device-relevant evidence rather than rely on general claims of Class III experience. Before transfer or supplier approval, confirm the proposed scope, validation responsibilities, traceability, change control, and record access. Suitability depends on the device and manufacturing work, not a broad capability statement alone.
Is ISO 13485 certification enough to qualify a Class III device manufacturer?
No. ISO 13485 is an important medical-device quality management standard, but certification alone does not establish suitability for every device or process. Assess the certificate’s scope and verify which sites and activities it covers. Also review relevant manufacturing capability, validation approach, traceability, risk controls, and supplier oversight. Qualification should account for the device’s intended use and regulatory pathway, as well as the specific evidence your program requires from the manufacturing partner.
What cleanroom class is needed to manufacture a Class III medical device?
There is no one cleanroom classification required for every Class III device. The appropriate environment depends on the device, materials, manufacturing steps, contamination risks, and applicable specifications, not on device class alone. Define environmental needs from the product and process requirements. Ask the manufacturer to explain relevant facility controls, monitoring, and validation, then verify that the proposed cleanroom scope suits the specific assembly or production work.
How should a company validate a Class III device manufacturing process?
Start by defining process requirements, risks, acceptance criteria, and responsibilities with the manufacturing partner. Use a documented validation plan suited to the process and its risks. It may include IQ to confirm equipment installation, OQ to assess operation within defined limits, and PQ to evaluate performance under intended production conditions. Retain protocols, results, deviations, approvals, and change records. Validation is not a generic checklist or a guarantee of regulatory acceptance.
What should buyers ask before selecting a Class III contract manufacturer?
Ask whether the supplier’s quality-system scope covers the proposed work, which processes and facilities fit the device, and what validation evidence is available. Clarify design and process ownership, traceability, change control, testing, packaging, and record access. Request relevant capability examples without expecting confidential customer information. Also establish how issues will be communicated and responsibilities documented. Resolving these points before production transfer helps both parties understand the scope and evidence expectations.